A New Direction at FDA? CTP Director Bret Koplow Talks Harm Reduction, Flavors and Faster PMTA Reviews
Posted by Alli Boughner on Sep 18th 2026
For years, the vapor industry has asked the FDA for something that should not be controversial: acknowledge the continuum of risk, give adult smokers accurate information about lower-risk alternatives, and create a regulatory pathway that actually allows innovative products to reach the market.
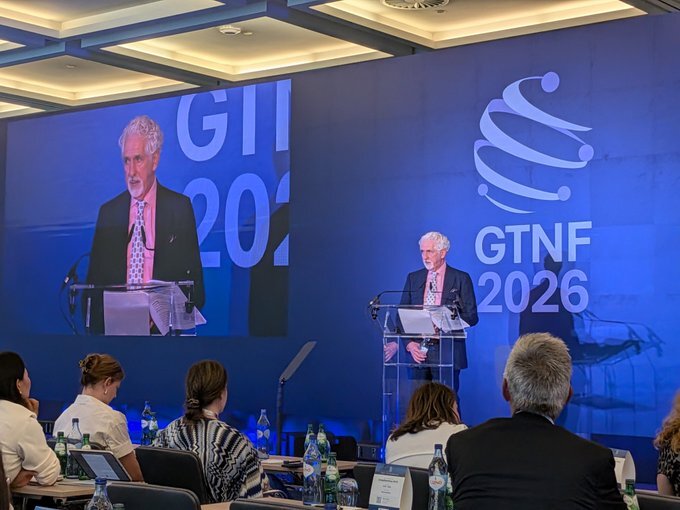
At the 2026 Global Tobacco & Nicotine Forum (GTNF) in Lisbon, newly appointed permanent Center for Tobacco Products (CTP) Director Bret Koplow signaled that those ideas may finally have a much larger role inside FDA.
Koplow outlined a vision for CTP that places tobacco harm reduction more squarely within the agency’s mission. His remarks covered everything from flavored vapor products and nicotine pouches to PMTA reform, youth access technology, physician education and the enormous market of unauthorized products.
For manufacturers, distributors and retailers, there was a lot to unpack.
Harm Reduction Is Finally Part of the Conversation
One of the most significant takeaways from Koplow’s remarks was his willingness to directly discuss differences in risk between nicotine products.
CTP’s priorities, he explained, include preventing youth initiation while also helping adults who smoke either quit completely or transition to products that present meaningfully lower health risks.
That distinction matters.
Combustible cigarettes are responsible for the overwhelming majority of tobacco-related disease and death, yet public understanding of relative risk has deteriorated significantly over the past decade.
Koplow pointed to data showing that fewer adults now correctly understand that FDA-authorized e-cigarettes are generally less harmful than smoking than did a decade ago.
That isn't merely an education problem. It can become a harm-reduction problem when adults who smoke are discouraged from switching because they incorrectly believe the alternatives are just as dangerous.
Koplow acknowledged that FDA has a role to play in correcting those misconceptions.
FDA Says It Does Not Have a Flavor Ban
Perhaps one of the most attention-grabbing moments came when Koplow addressed flavored vapor products.
His message was straightforward: FDA does not have a blanket flavor ban.
Instead, he said the agency's concern is the potential for youth use. That distinction opens an important door for manufacturers capable of demonstrating that youth-access risks can be sufficiently mitigated.
We have already seen an example of this approach.
Earlier this year, FDA authorized Glas vapor products in non-tobacco and non-menthol flavors using technology designed to restrict youth access. Those protections included identity verification and other technological controls.
At GTNF, Koplow suggested that this may only be the beginning.
If manufacturers can demonstrate that technology meaningfully limits youth access, FDA could potentially become more receptive to a broader range of flavored products intended for adults who smoke.
That does not mean flavored products suddenly have an easy path through the PMTA process. Applicants still have to satisfy the statutory “appropriate for the protection of public health” standard and demonstrate that the expected benefits to adults outweigh potential population-level risks.
But after years in which flavored vapor applications faced extraordinary regulatory barriers, even acknowledging that flavors themselves are not automatically disqualifying is significant.
FDA Is Testing a Faster PMTA Pathway
Koplow also revealed that CTP is testing a supplemental PMTA process for certain modifications to products that have already received marketing authorization.
The concept makes sense.
If FDA has already reviewed a product's manufacturing processes, scientific evidence and other foundational information, a manufacturer making a relatively minor modification should not necessarily have to recreate an entirely new application from the ground up.
Under the pilot, FDA may be able to rely on information it has already reviewed and focus on what actually changed.
If successful, this could reduce unnecessary duplication and allow improvements to authorized products to reach consumers more quickly.
The initiative also reflects a broader change Koplow repeatedly emphasized: CTP is moving away from treating every tobacco product category exactly the same.
Lessons From the Nicotine Pouch Pilot
FDA has already been experimenting with that approach through its nicotine pouch review pilot.
Rather than automatically putting every application through the same review structure, CTP has been examining whether the scope of scientific review can be tailored to the actual risk profile of a product category.
According to Koplow, the first authorization produced through the pilot was completed in less than 180 days — an important milestone considering the years-long PMTA delays that have plagued the agency.
The pilot has not been perfect. Some applications were less complete than FDA anticipated, slowing the process.
But CTP appears to be taking what it has learned and applying those lessons elsewhere.
That could be important for vapor manufacturers that have spent years arguing that applying a one-size-fits-all regulatory framework to dramatically different tobacco and nicotine products makes little scientific sense.
An Important Statement About Nicotine Pouches
Koplow also made a particularly notable observation about nicotine pouches, describing them as generally representing the lowest-risk category among tobacco products.
He raised another issue that receives relatively little attention: nicotine delivery.
Adults accustomed to cigarettes or other products that deliver nicotine quickly may initially find pouches less satisfying because nicotine delivery can be slower. If consumers don't understand that difference, they may abandon the lower-risk product and return to smoking.
For a regulator focused on population-level health, that matters.
Successful harm reduction isn't simply about making alternatives technically available. Adults who smoke also need accurate information about what to expect when switching.
FDA Admits It Needs to Do a Better Job Explaining the PMTA Process
The industry has heard plenty from FDA about inadequate PMTA applications.
At GTNF, Koplow acknowledged another side of the problem: FDA itself needs to communicate its expectations better.
CTP says more than 99% of the more than two million applications it received lacked basic information necessary for review. Many vapor businesses submitting applications were small manufacturers without the scientific and regulatory infrastructure of major tobacco companies.
Applications frequently lacked evidence FDA expected involving product characterization, manufacturing consistency, pharmacokinetics or potential consumer benefits.
But Koplow acknowledged that FDA bears some responsibility for helping applicants understand what the agency expects.
CTP is now developing additional guidance on common application deficiencies and encouraging manufacturers to meet with the agency before conducting expensive studies that may ultimately fail to answer FDA's questions.
That kind of communication should have been standard practice from the beginning.
Manufacturers cannot reasonably be expected to navigate a regulatory process when the standards become clear only after applications have already been submitted — or denied.
The PMTA Backlog Is Shrinking
There are also signs that CTP's review process is moving faster.
FDA previously reported reducing its application backlog by approximately 70% during 2025.
Koplow said the initial stages of PMTA review have also accelerated considerably. Acceptance review is now generally taking between one and three weeks, while filing review takes roughly 21 days.
Those are improvements.
However, staffing remains a major obstacle. CTP does not have enough scientific review teams to process applications at the pace leadership would like, and Koplow said the center would like to more than double that capacity.
So while progress is being made, the fundamental problem of getting products through full scientific review and to a final decision has not disappeared.
The Unauthorized Market Remains a Major Problem
Faster authorization also matters because manufacturers attempting to comply with FDA requirements are competing against a massive unauthorized marketplace.
Koplow acknowledged the obvious fairness problem this creates.
Companies can invest millions of dollars in scientific studies, manufacturing controls and PMTA submissions while competitors sell products that have never undergone FDA review.
Meanwhile, consumers have very little ability to distinguish between the two.
There is currently no simple indicator on most packaging telling consumers whether FDA has actually reviewed and authorized a particular product.
CTP is reportedly exploring ways to make that distinction clearer.
FDA and Customs have also dramatically increased import enforcement. Koplow said import refusals grew from roughly 100 in fiscal year 2023 to more than 9,000 in fiscal year 2025.
Yet unauthorized products continue to enter and remain on the U.S. market.
For manufacturers actually investing in compliance, enforcement and regulatory efficiency have to improve together. Speeding up PMTA reviews accomplishes little if companies that ignore the process continue enjoying a competitive advantage.
Could This Be a Different CTP?
That may ultimately be the biggest takeaway from GTNF.
None of these announcements magically fixes the tobacco regulatory system.
There are still enormous questions surrounding PMTA timelines, flavored-product standards, enforcement, state registry laws and whether independent manufacturers will finally have a realistic opportunity to obtain marketing authorization.
But the language coming from CTP leadership is noticeably different.
FDA's tobacco center is now publicly discussing tobacco harm reduction, the continuum of risk, correcting misinformation about nicotine, improving adult access to lower-risk products and making the authorization process more efficient.
For an industry that has spent years asking regulators to acknowledge those very principles, that deserves attention.
The real test, however, will be what happens next.
Manufacturers don't need another shift in rhetoric. They need clear guidance, predictable standards, timely decisions and a regulatory pathway that gives scientifically supported products — including those developed by independent companies — a genuine opportunity to reach adult consumers.
If Koplow's vision translates into those kinds of changes, GTNF 2026 may eventually be remembered as more than another industry conference.
It may mark the beginning of a very different chapter at FDA's Center for Tobacco Products.